


 癌症的基本特征包括细胞增殖、血管生成、迁移、凋亡逃避机制和细胞永生等。找到癌症发生过程中这些通路的关键标记物和对应的抗体用于检测至关重要。
 为您推荐一个泛素化位点预测神器——泛素化分析工具,可以为您的蛋白的泛素化位点作出预测和评分。
 细胞自噬受体图形绘图工具为你的蛋白的细胞受体结合位点作出预测和评分,识别结合到自噬通路中的蛋白是非常重要的,便于让我们理解自噬在正常生理、病理过程中的作用,如发育、细胞分化、神经退化性疾病、压力条件下、感染和癌症。
TTC21B Rabbit pAb
TTC21B Rabbit pAb
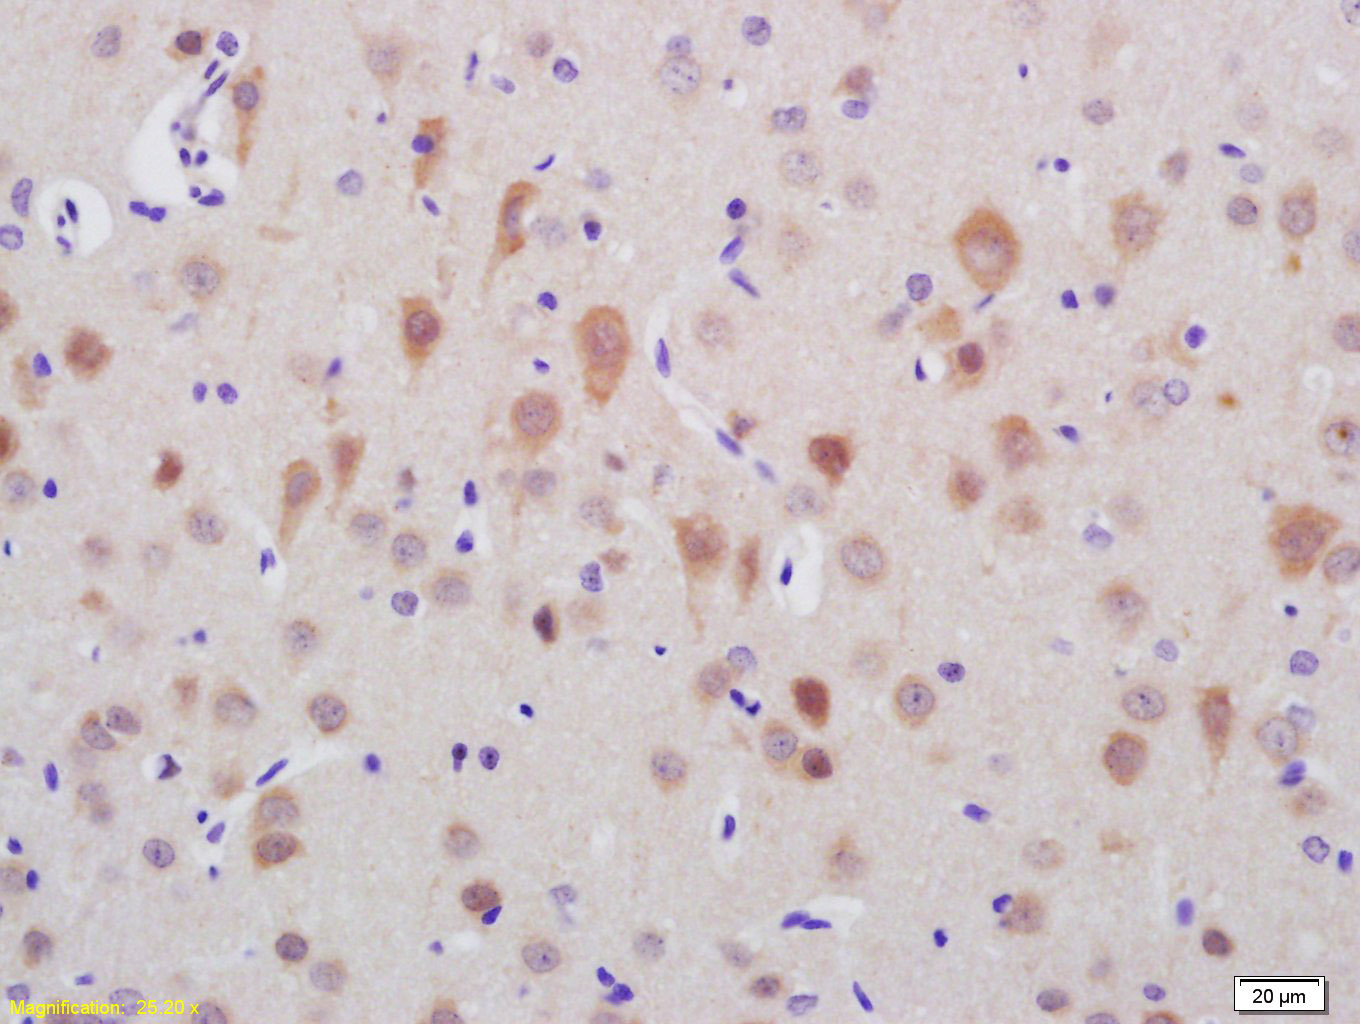
- 产品详情
- 实验流程
- 背景知识
Application 
| IHC-P, IHC-F, IF |
|---|---|
| Primary Accession | Q7Z4L5 |
| Reactivity | Rat |
| Predicted | Pig, Dog, Human, Mouse, Rabbit, Horse, Sheep |
| Host | Rabbit |
| Clonality | Polyclonal |
| Calculated MW | 150937 Da |
| Physical State | Liquid |
| Immunogen | KLH conjugated synthetic peptide derived from human TTC21B |
| Epitope Specificity | 1221-1316/1316 |
| Isotype | IgG |
| Purity | affinity purified by Protein A |
| Buffer | 0.01M TBS (pH7.4) with 1% BSA, 0.02% Proclin300 and 50% Glycerol. |
| SUBCELLULAR LOCATION | Cytoplasm. |
| DISEASE | Note=Ciliary dysfunction leads to a broad spectrum of disorders, collectively termed ciliopathies. Overlapping clinical features include retinal degeneration, renal cystic disease, skeletal abnormalities, fibrosis of various organ, and a complex range of anatomical and functional defects of the central and peripheral nervous system. The ciliopathy range of diseases includes Meckel-Gruber syndrome, Bardet-Biedl syndrome, Joubert syndrome, nephronophtisis, Senior-Loken syndrome, and Jeune asphyxiating thoracic dystrophy among others. TTC21B is causally associated with diverse ciliopathies, and also acts as a modifier gene across the ciliopathy spectrum. TTC21B mutations interact in trans with mutations in other ciliopathy-causing genes and contribute to disease manifestation and severity. Defects in TTC21B are the cause of nephronophthisis type 12 (NPHP12) [MIM:613820]. NPHP12 is an autosomal recessive disorder resulting in end-stage renal disease. It is a progressive tubulo-interstitial kidney disorder histologically characterized by modifications of the tubules with thickening of the basement membrane, interstitial fibrosis and, in the advanced stages, medullary cysts. Some patients manifest extra-renal features including retinal, skeletal and central nervous system defects. Defects in TTC21B are the cause of asphyxiating thoracic dystrophy type 4 (ATD4) [MIM:613819]. ATD4 is an autosomal recessive chondrodysplasia characterized by a severely constricted thoracic cage, short-limbed short stature, and polydactyly. It often leads to death in infancy because of respiratory insufficiency. Retinal degeneration, cystic renal disease and hepatic disease can be present in affected individuals who survive early childhood. Defects in TTC21B may be a cause of Bardet-Biedl syndrome (BBS) [MIM:209900]. A syndrome characterized by usually severe pigmentary retinopathy, early-onset obesity, polydactyly, hypogenitalism, renal malformation and mental retardation. Secondary features include diabetes mellitus, hypertension and congenital heart disease. Bardet-Biedl syndrome inheritance is autosomal recessive, but three mutated alleles (two at one locus, and a third at a second locus) may be required for clinical manifestation of some forms of the disease. Defects in TTC21B may be a cause of Joubert syndrome (JBTS) [MIM:213300]. A disorder presenting with cerebellar ataxia, oculomotor apraxia, hypotonia, neonatal breathing abnormalities and psychomotor delay. Neuroradiologically, it is characterized by cerebellar vermian hypoplasia/aplasia, thickened and reoriented superior cerebellar peduncles, and an abnormally large interpeduncular fossa, giving the appearance of a molar tooth on transaxial slices (molar tooth sign). Additional variable features include retinal dystrophy and renal disease. |
| Important Note | This product as supplied is intended for research use only, not for use in human, therapeutic or diagnostic applications. |
| Background Descriptions | This gene encodes a member of TTC21 family, containing several tetratricopeptide repeat (TPR) domains. This protein is localized to the cilium axoneme, and may play a role in retrograde intraflagellar transport in cilia. Mutations in this gene are associated with various ciliopathies, nephronophthisis 12, and asphyxiating thoracic dystrophy 4. [provided by RefSeq, Oct 2011] |
| Gene ID | 79809 |
|---|---|
| Other Names | Tetratricopeptide repeat protein 21B, TPR repeat protein 21B, Intraflagellar transport 139 homolog, TTC21B (HGNC:25660) |
| Dilution | IHC-P=1:100-500,IHC-F=1:100-500,IF=1:100-500 |
| Storage | Store at -20 °C for one year. Avoid repeated freeze/thaw cycles. When reconstituted in sterile pH 7.4 0.01M PBS or diluent of antibody the antibody is stable for at least two weeks at 2-4 °C. |
For Research Use Only. Not For Use In Diagnostic Procedures.
| Name | TTC21B (HGNC:25660) |
|---|---|
| Function | Component of the IFT complex A (IFT-A), a complex required for retrograde ciliary transport and entry into cilia of G protein- coupled receptors (GPCRs). Essential for retrograde trafficking of IFT- 1, IFT-B and GPCRs (PubMed:27932497). Negatively modulates the SHH signal transduction (By similarity). |
| Cellular Location | Cytoplasm, cytoskeleton, cilium axoneme {ECO:0000250|UniProtKB:Q0HA38} |
Research Areas
Application Protocols
Provided below are standard protocols that you may find useful for product applications.
BACKGROUND
This product as supplied is intended for research use only, not for use in human, therapeutic or diagnostic applications.

终于等到您。ABCEPTA(百远生物)抗体产品。
点击下方“我要评价 ”按钮提交您的反馈信息,您的反馈和评价是我们最宝贵的财富之一,
我们将在1-3个工作日内处理您的反馈信息。
如有疑问,联系:0512-88856768 tech-china@abcepta.com.


