


 癌症的基本特征包括细胞增殖、血管生成、迁移、凋亡逃避机制和细胞永生等。找到癌症发生过程中这些通路的关键标记物和对应的抗体用于检测至关重要。
 为您推荐一个泛素化位点预测神器——泛素化分析工具,可以为您的蛋白的泛素化位点作出预测和评分。
 细胞自噬受体图形绘图工具为你的蛋白的细胞受体结合位点作出预测和评分,识别结合到自噬通路中的蛋白是非常重要的,便于让我们理解自噬在正常生理、病理过程中的作用,如发育、细胞分化、神经退化性疾病、压力条件下、感染和癌症。
NAGA Rabbit pAb
NAGA Rabbit pAb
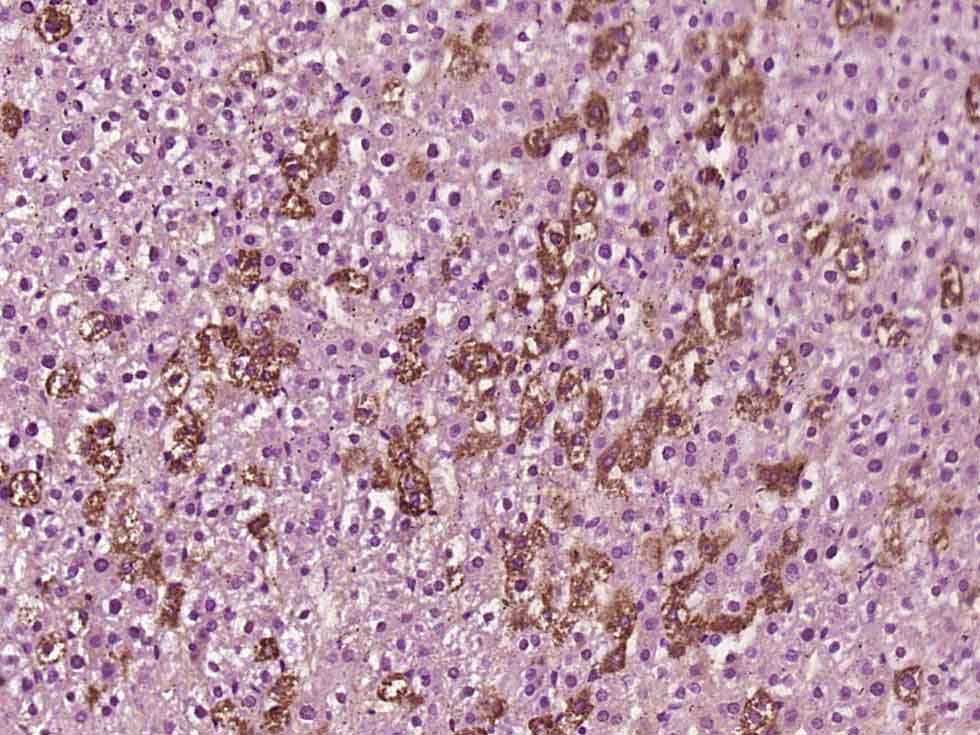
- 产品详情
- 实验流程
- 背景知识
Application 
| IHC-P, IHC-F, IF |
|---|---|
| Primary Accession | P17050 |
| Reactivity | Rat |
| Predicted | Pig, Dog, Human, Mouse, Rabbit, Horse, Sheep |
| Host | Rabbit |
| Clonality | Polyclonal |
| Calculated MW | 46565 Da |
| Physical State | Liquid |
| Immunogen | KLH conjugated synthetic peptide derived from human NAGA |
| Epitope Specificity | 101-200/411 |
| Isotype | IgG |
| Purity | affinity purified by Protein A |
| Buffer | 0.01M TBS (pH7.4) with 1% BSA, 0.02% Proclin300 and 50% Glycerol. |
| SUBCELLULAR LOCATION | Lysosome. |
| DISEASE | Schindler disease (SCHIND) [MIM:609241]: Form of NAGA deficiency characterized by early-onset neuroaxonal dystrophy and neurological signs (convulsion during fever, epilepsy, psychomotor retardation and hypotonia). NAGA deficiency is typically classified in three main phenotypes: NAGA deficiency type I (Schindler disease or Schindler disease type I) with severe manifestations; NAGA deficiency type II (Kanzazi disease or Schindler disease type II) which is mild; NAGA deficiency type III (Schindler disease type III) characterized by mild-to-moderate neurologic manifestations. NAGA deficiency results in the increased urinary excretion of glycopeptides and oligosaccharides containing alpha-N-acetylgalactosaminyl moieties. Inheritance is autosomal recessive. Note: The disease is caused by mutations affecting the gene represented in this entry. Ref.13 Ref.15 Kanzaki disease (KANZD) [MIM:609242]: Autosomal recessive disorder characterized by late-onset, angiokeratoma corporis diffusum and mild intellectual impairment. Note: The disease is caused by mutations affecting the gene represented in this entry. |
| Important Note | This product as supplied is intended for research use only, not for use in human, therapeutic or diagnostic applications. |
| Background Descriptions | NAGA encodes the lysosomal enzyme alpha-N-acetylgalactosaminidase, which cleaves alpha-N-acetylgalactosaminyl moieties from glycoconjugates. Mutations in NAGA have been identified as the cause of Schindler disease types I and II (type II also known as Kanzaki disease). [provided by RefSeq, Jul 2008] |
| Gene ID | 4668 |
|---|---|
| Other Names | Alpha-N-acetylgalactosaminidase, 3.2.1.49, Alpha-galactosidase B, NAGA (HGNC:7631) |
| Dilution | IHC-P=1:100-500,IHC-F=1:100-500,IF=1:100-500 |
| Storage | Store at -20 °C for one year. Avoid repeated freeze/thaw cycles. When reconstituted in sterile pH 7.4 0.01M PBS or diluent of antibody the antibody is stable for at least two weeks at 2-4 °C. |
For Research Use Only. Not For Use In Diagnostic Procedures.
| Name | NAGA (HGNC:7631) |
|---|---|
| Function | Removes terminal alpha-N-acetylgalactosamine residues from glycolipids and glycopeptides. Required for the breakdown of glycolipids. |
| Cellular Location | Lysosome. |
Research Areas
Application Protocols
Provided below are standard protocols that you may find useful for product applications.
BACKGROUND
This product as supplied is intended for research use only, not for use in human, therapeutic or diagnostic applications.

终于等到您。ABCEPTA(百远生物)抗体产品。
点击下方“我要评价 ”按钮提交您的反馈信息,您的反馈和评价是我们最宝贵的财富之一,
我们将在1-3个工作日内处理您的反馈信息。
如有疑问,联系:0512-88856768 tech-china@abcepta.com.


